

FDA
Latest News
Advertisement
Latest Videos
Shorts
0:28
DIA 2026: FDA's New Stance, Quality by Design in Clinical Trials
4 months ago
by
Kevin Bugin(+1 more)
0:51
Bringing Together Modern Infrastructure to Meet New FDA Standards
5 months ago
by
Raj Indupuri(+1 more)
1:15
ACT Ops Take: Understanding the FDA's CNPV Program and Review Timelines
5 months ago
by
Andy Studna, Senior Editor
0:51
A Major Shift in FDA Approval for Rare Disease Therapies
6 months ago
by
Mwango Kashoki, MD, MPH(+1 more)
0:59
ACT Ops Take: FDA Actions Reshaping Drug Development in 2026
7 months ago
by
Andy Studna, Senior Editor
0:48
The Shift from Frequentist to Bayesian Mindsets
8 months ago
by
David Morton, PhD(+1 more)
1:30
ACT Ops Take: How Bayesian Methods Revolutionize Rare Disease Trials
8 months ago
by
Andy Studna, Senior Editor
0:46
How FDA Uncertainty is Changing Regulatory Strategy
9 months ago
by
Charlie Paterson(+1 more)
Advertisement
More News
Advertisement
Real-time RBQM requires proportionate analysis frequency matched to data arrival and issue criticality, not continuous reanalysis, because signal churn from over-frequent runs creates alert fatigue and operational noise that obscures meaningful deterioration.
A new FDA Voices post from the agency's acting chief of staff details completed milestones and upcoming priorities across complete response letter disclosure, application lifecycle transparency, new approach methodologies, labeling, and inspectional records.
A new request for information asks for comments on dose selection, safety monitoring, eligibility criteria, and oversight approaches to support the responsible development of ibogaine drug products.
In rare disease drug development where no registry or natural history dataset exists, real-world evidence quality depends on treating evidence engineering as a design-stage decision, defining intended regulatory use and standardizing endpoints, harmonization, and data provenance upfront rather than reconciling gaps after collection.
FDA Opens Applications for Expedited IND Pilot Designed to Accelerate First-in-Human Trial Timelines
The pilot pairs drug sponsors with qualified research institutions to compress the path from drug identification to first-in-human study through rolling submission review and earlier coordination of institutional review board and site activation activities.
Senior FDA leaders outline concrete steps to strengthen inspection coverage and data integrity review as the globalization of clinical research strains the agency's oversight capacity.
In this episode of Beyond Compliance, Otis Johnson, PhD, MPA, founder and principal consultant at Vantix Operations, speaks with Joel Stanley, CEO of AJNA BioSciences, about why botanical therapies need pharmaceutical-grade standardization and clinical evidence to earn physician trust and insurance coverage.
A new discussion paper from the FDA's Center for Devices and Radiological Health proposes a two-axis risk framework and competency-based evaluation approach for generative artificial intelligence-enabled medical devices.
In this Q&A, Raviv Pryluk, PhD, CEO and co-founder of PhaseV, discusses the data standardization bottleneck that sits between raw trial data and real-time analysis, what the FDA's continuous monitoring pilot will require to scale beyond large pharma, and why an ecosystem approach is the only path to making real-time trial oversight a practical reality.
A new survey of 37 US biotechnology executives finds broad preference for conducting first-in-human trials domestically, with regulatory unpredictability and clinical holds cited as the primary drivers of overseas migration.
Advertisement
From point-of-care sample collection to real-world data curation, wearable device integration, and continuous review infrastructure, data quality is not a single problem in clinical trials—it is a layered challenge that manifests differently at every stage of development and demands a different set of solutions at each one.
In this Q&A, Richard Graham, PhD, co-founder and chairman of the board at TruTechnologies, discusses why six decades of manual site-level processes continue to undermine clinical trial execution, what national initiatives like Operation TrialBlazer leave unaddressed, and why meaningful modernization has to start with data collected at the point of care.
Acting CBER director Karim Mikhail outlines the structural barriers slowing Phase I trials in the US and details how the FDA's Expedited IND Pilot is designed to address them.
Real-time AI-enabled decision-making only accelerates development when trials have coordinated operational infrastructure for randomization, supply management, and workflows capable of responding to new insights without disrupting execution or compliance.
As agentic AI, automated data harmonization, and real-time monitoring reshape clinical development, the organizations seeing meaningful results are those that have invested in unified data infrastructure and disciplined governance rather than cycling through pilots without the foundation to scale them.
New documents targeting performance status, washout periods, and laboratory value thresholds aim to close the gap between patients willing to participate in oncology trials and those who actually enroll.
In this Q&A, Gaynor Anders, chief delivery officer at Trialbee, discusses how the FDA's single pivotal trial shift is raising the stakes on patient recruitment, why the study-by-study model has persisted for so long, and what program-level recruitment actually requires from sponsors in terms of infrastructure, data sharing, and organizational change.
The agency's guidance addresses the unique challenges of studying compounds like psilocybin and MDMA, from functional unblinding to safety monitoring and abuse potential assessment.
In this Q&A, Rob DiCicco, vice president of portfolio management at TransCelerate Biopharma, discusses the collaborative work underway with FDA and CTTI to explore selective safety data collection as a model for de-risking trial design, reducing site and patient burden, and scaling pragmatic trial approaches across the industry.
The FDA's launch of real-time clinical trial proof-of-concept studies signals a fundamental shift in regulatory oversight, one that most sponsors are not yet equipped to meet and that demands urgent investment in unified data infrastructure, quality-by-design practices, and protocol digitization.
Clinical data security built into system architecture from the foundation, rather than layered on afterward, reduces regulatory risk and actually accelerates research by removing manual compliance burden and improving team confidence in data integrity.
In this Q&A, Raj Indupuri, CEO and co-founder of eClinical Solutions, discusses what the FDA's push toward continuous data review actually demands of sponsors operationally, why fragmented systems are the core obstacle, and how AI and real-world evidence fit into a more data-driven regulatory environment.
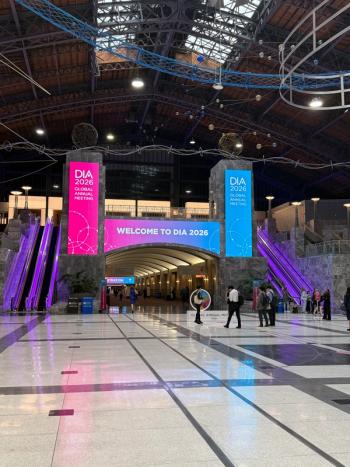
From real-time evidence generation to federated AI to site-level data integration, ACT spoke with seven experts at DIA 2026 on the trends and challenges defining clinical trial operations today.
A coordinated department-wide initiative combines FDA regulatory reforms, NIH investments, and new agency programs to reduce development timelines and reverse a growing shift of clinical research overseas.
From rising costs and regulatory uncertainty to persistent vaccine hesitancy, sponsors face mounting pressure to standardize operations, build community trust, and develop the behavioral capabilities needed to run vaccine trials effectively in a rapidly shifting environment.
Advertisement
Advertisement
Trending on Applied Clinical Trials Online
1
How Much of the Community Participation Gap Is About Infrastructure, Not Willingness?
2
Closing the Medical Writing Capacity Gap: Why Training Beats Technology
3
Measuring What Matters: Realigning Clinical Endpoints with Patient Reality
4
Clinical Trials Do Not Have a Data Problem, They Have a Decision-Readiness Problem
5