


FDA
Latest News
Advertisement
Latest Videos
Advertisement
More News
Advertisement
In this video interview, Mwango Kashoki, MD, MPH, senior vice president and global head of regulatory strategy at Parexel, outlines the steps sponsors can take today—from early FDA engagement and robust non-clinical programs to adaptive trial designs and confirmatory evidence packages—to avoid delays as the guidance moves toward finalization.
In this video interview, Mwango Kashoki, MD, MPH, senior vice president and global head of regulatory strategy at Parexel, discusses how heterogeneity in disease course and patient characteristics creates confounding risk when using natural history data as an external control, and what sponsors need to do to achieve meaningful patient matching.
Agency outreach targets a compliance gap affecting thousands of registered trials, with nearly 30% of studies subject to mandatory reporting showing no results submitted to ClinicalTrials.gov.
In part 3 of this three-part series, behavioral science reveals that clinical trial diversity cannot be achieved through last-minute recruitment efforts, but requires designing protocols, sites, and enrollment strategies around the real barriers and needs of underrepresented communities from the outset.
Moving beyond vendor evaluation to incentive design in Phase II/III rare and genetic disease trials.
In the first months of 2026, the FDA has moved on multiple fronts to modernize how drugs are developed, evaluated, and approved, signaling a broader regulatory shift toward flexibility, mechanism-based evidence, and human-centric science.
New framework establishes scientific standards for alternative methodologies, signaling a regulatory shift toward human-centric data in preclinical drug development.
Draft guidance outlines a “plausible mechanism” pathway to support approval of highly targeted therapies when traditional trials are not feasible.
As rare disease trials face persistent feasibility challenges, Bayesian designs are gaining momentum by enabling more flexible, data-driven approaches that integrate prior knowledge, reduce sample size demands, and align with evolving FDA guidance to accelerate evidence generation.
Following a Type A meeting with the FDA’s Center for Biologics Evaluation and Research, Moderna’s flu vaccine candidate mRNA-1010 has been accepted for formal review after the company revised its regulatory strategy.
Advertisement
FDA is shifting its default standard for drug approvals, announcing that one adequate and well-controlled clinical trial will generally be sufficient for approvals.
Charlie Paterson, partner at PA Consulting, discusses how fewer new guidance updates are pushing sponsors to rely on historical precedents and non-US standards when making trial design and operational decisions.
The FDA and EMA have aligned on ten guiding principles for the responsible use of artificial intelligence across the drug development lifecycle, establishing a shared framework to support innovation, regulatory consistency, and patient safety.
Charlie Paterson, partner at PA Consulting, explains how reduced FDA capacity and staff turnover have led to longer regulatory timelines, increased preparation for agency interactions, and delayed feedback during early trial planning.
A look at how efficiency, access, platformization, AI, non-traditional players, and regulatory recovery are expected to reshape clinical operations in 2026.
Examine how the FDA’s acceptance of de-identified real-world evidence shifts clinical operations workflows and why understanding the difference between pseudonymized and anonymized data is now critical for privacy, compliance, and evidence generation.
The FDA’s new draft guidance on Bayesian methodology signals a shift toward more flexible, data-driven clinical trial designs, enabling sponsors to use prior data and adaptive approaches to improve efficiency, reduce timelines, and support regulatory decision-making.
The clinical trial ecosystem is entering a phase of consolidation and reinvention driven by the collapse of boundaries between functions, data, and even companies themselves.
What Patients Want From Remote Trials: New Survey Data Reveals Preferences Around Usability, Trust, and Participation
ByJiang Li
New survey data show a strong patient preference for fully remote clinical trials, underscoring how convenience, intuitive technology, and FDA-cleared digital tools are reshaping enrollment and retention strategies as decentralized models become a lasting fixture in clinical research.
New FDA guidance signals a major shift for CAR-T development, calling for randomized trials with standard-of-care control groups and clear evidence of superiority over existing therapies, while simultaneously easing REMS requirements to reduce logistical burdens for treatment centers and patients.
Former FDA Commissioners Warn New Vaccine Policies Could Undermine Longstanding Regulatory Framework
Twelve former FDA leaders have publicly challenged the agency’s proposed overhaul of vaccine approvals, sparked by an internal memo linking child deaths to COVID-19 vaccination, arguing the changes threaten evidence-based standards, weaken immunobridging practices, and risk eroding public trust.
The FDA has introduced a secure, agency-wide agentic AI system to support complex regulatory workflows—enhancing reviews, surveillance, and inspections—while maintaining strict data protections and human oversight.
In this Q&A, John Kirk, principal regulatory strategist at Veristat, explains how the FDA’s Commissioner’s National Priority Voucher (CNPV) Program is accelerating operational timelines, tightening cross-functional workflows, and redefining what sponsors and CROs must deliver to secure and execute an ultra-compressed one-to-two-month review.
The company submitted the new dosage for approval based on its Phase III trial results.
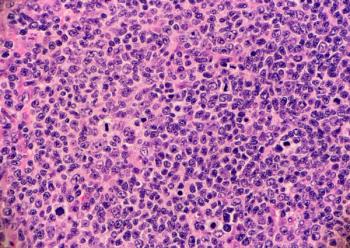
Epkinly plus rituximab and lenalidomide is the first bispecific antibody combination FDA-approved for relapsed or refractory follicular lymphoma, backed by Phase III data showing substantially improved disease control over standard therapy.
Advertisement
Advertisement
Trending on Applied Clinical Trials Online
1
The Control Plane for Agentic Clinical Operations: Governing Autonomous Digital Actors in Regulated Trials
2
HHS Launches SURPASS Program and Three Complementary Initiatives to Transform Clinical Trial Design and Infrastructure
3
Hierarchical Endpoints in Cardiology: How Much Treatment Benefit, and From Where
4
What ClinOps Professionals Just Starting With AI Should Keep Top of Mind
5